Mn2O4 computation [ESPRESSO]

This page presents an overview of the calculation performed for Mn2O4.

NOTE: This page is refreshed every 3 min as results are produced. Reload manually (Ctrl-R) to force update.

The following computations are requested:

The calculator being used is: ESPRESSO which detailed configuration follows:

Calculator Configuration{'template': , 'profile': , 'directory': PosixPath('/var/lib/material-modelling/data/MnO2-Pyrolusite.cif-QuantumEspresso-farhie-20261101-190957'), 'parameters': {'input_data': {'control': {'outdir': '/var/lib/material-modelling/data/MnO2-Pyrolusite.cif-QuantumEspresso-farhie-20261101-190957', 'prefix': 'Mn2O4', 'verbosity': 'high', 'tstress': True, 'tprnfor': True, 'disk_io': 'low'}, 'system': {'ecutwfc': 30.0, 'occupations': 'smearing', 'nosym': False, 'noinv': True, 'input_dft': 'PBE', 'smearing': 'mv', 'degauss': 0.009554823852044876, 'lda_plus_u': True, 'lda_plus_u_kind': 0, 'U_projection_type': 'atomic', 'Hubbard_U(1)': 3.9}, 'electrons': {'conv_thr': 7.349864501572982e-07, 'electron_maxstep': 100, 'diagonalization': 'david', 'mixing_beta': 0.7}}, 'kpts': (np.int64(3), np.int64(3), np.int64(3)), 'pseudopotentials': {'Mn': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn1': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn2': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn3': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn4': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn5': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn6': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn7': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn8': 'mn_pbe_v1.5.uspp.F.UPF', 'Mn9': 'mn_pbe_v1.5.uspp.F.UPF', 'O': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O1': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O2': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O3': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O4': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O5': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O6': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O7': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O8': 'O.pbe-n-kjpaw_psl.0.1.UPF', 'O9': 'O.pbe-n-kjpaw_psl.0.1.UPF'}}, 'atoms': Atoms(symbols='Mn2O4', pbc=True, cell=[4.396, 4.396, 2.871], initial_magmoms=..., spacegroup_kinds=..., calculator=Espresso(...)), 'results': {}, 'use_cache': True}

The setting used are:

Settings{'version': '26.10.31', 'atoms': '/var/lib/material-modelling/data/MnO2-Pyrolusite.cif-QuantumEspresso-farhie-20261101-190957/MnO2-Pyrolusite.cif', 'calculator': 'ESPRESSO', 'compute': 'ELECTRONS_ELECTRONS_3D_PHONONS_PHONONS_3D_DIFFRACTION_FLUORESCENCE_XAS', 'convergence': 1e-05, 'diffraction_B': 0, 'diffraction_bins': 10000, 'diffraction_qmax': 25, 'directory': '/var/lib/material-modelling/data/MnO2-Pyrolusite.cif-QuantumEspresso-farhie-20261101-190957', 'displacement': 0.01, 'ecut': 408.17079, 'eigensolver': 'dav', 'epoints': 101, 'hubbard': 'auto', 'kgamma': None, 'kpoints3d': 11, 'kpoints': array([3, 3, 3]), 'label': 'Mn2O4', 'log': None, 'maxiter': None, 'mpi': 10, 'mpirun': '/usr/bin/mpirun -np 10', 'nbands': None, 'occupations': 0.13, 'parallel_displacements': 1, 'potentials': '/usr/share/espresso/pseudo', 'qgamma': None, 'qpoints3d': [-1, 1, 21], 'spin': 'auto', 'start_date': 'Sun Nov 1 19:09:58 2026', 'supercell': array([2, 2, 2]), 'temperature': 300, 'to_all': 'True', 'to_hdf5': True, 'to_html': True, 'to_json': True, 'tolerance': 1e-05, 'to_mat': True, 'to_mccode': True, 'to_npy': True, 'to_nrrd': True, 'to_png': True, 'to_svg': True, 'to_toml': True, 'wpoints': 51, 'xas_bins': 10000, 'xas_emax': None, 'xas_emin': None, 'xc': 'PBE', 'xray_energy': 38, 'xray_polar': array([1, 0, 0]), 'smearing': 'semiconductor', 'user_id': 'farhie', 'user_ip': ' [195.221.10.47]', 'chemical_symbols': ['Mn', 'Mn', 'O', 'O', 'O', 'O'], 'chemical_formula': 'Mn2O4', 'cell_parameters': array([ 4.396, 4.396, 2.871, 90. , 90. , 90. ]), 'kpts_density': np.int64(1296), 'Atoms': Atoms(symbols='Mn2O4', pbc=True, cell=[4.396, 4.396, 2.871], initial_magmoms=..., spacegroup_kinds=..., calculator=Espresso(...)), '_hubbard_printed': True, 'hubbard_used': {'Mn': 3.9}}

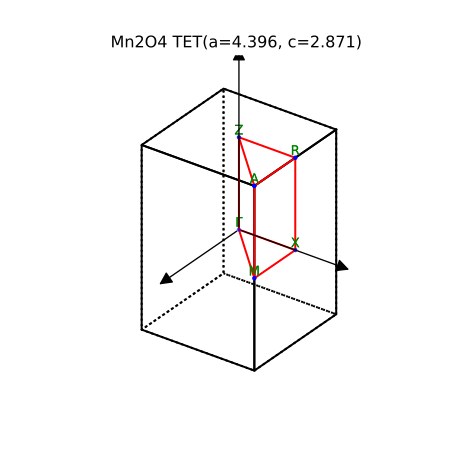
Lattice cell on Sun Nov 1 19:09:59 2026

Generated:

Ground state on Sun Nov 1 19:10:15 2026

Generated:

Properties:

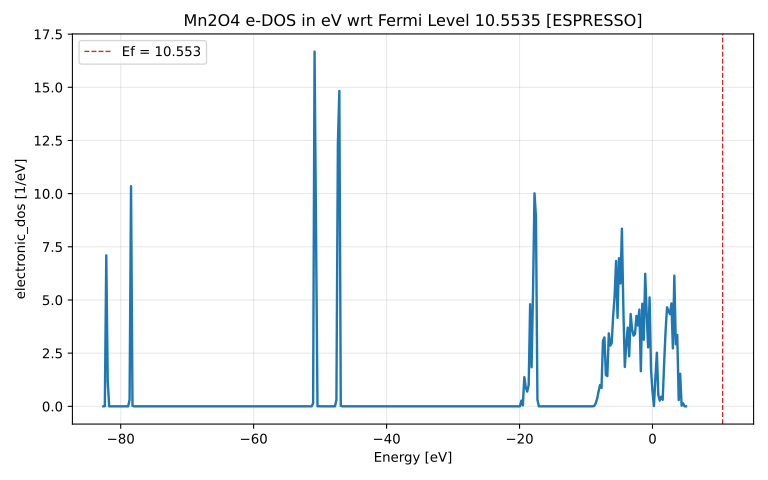
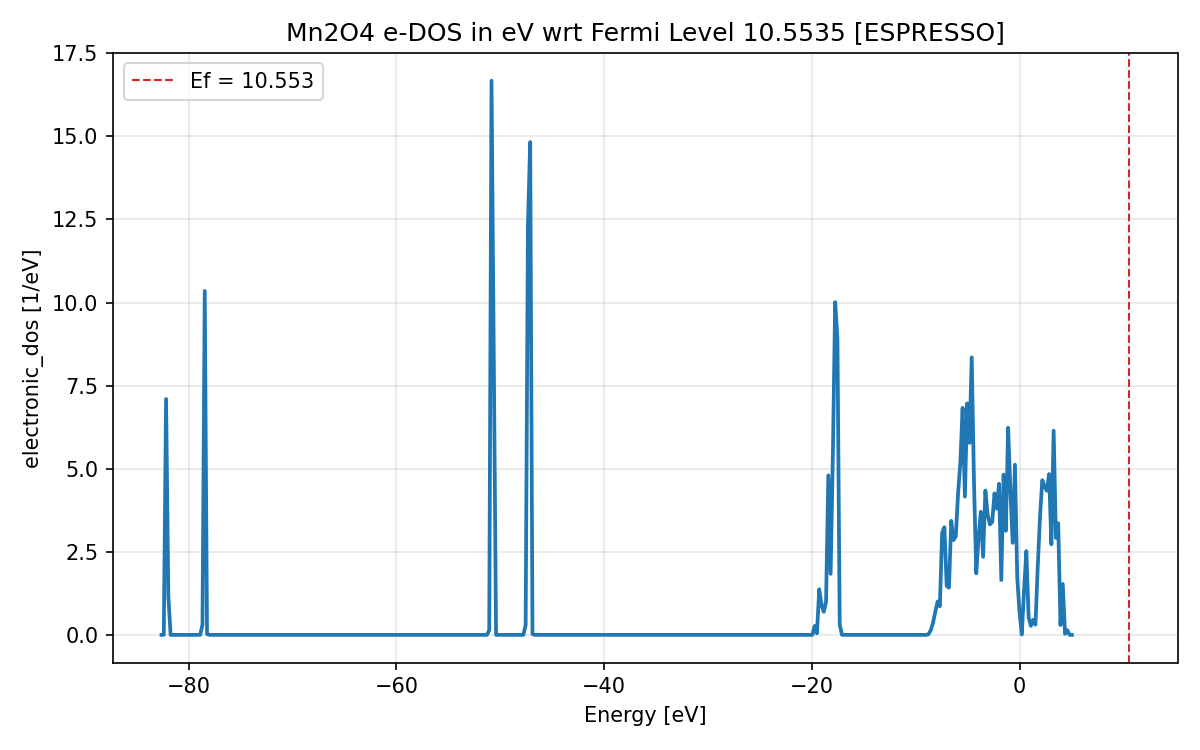
Electronic density of states on Sun Nov 1 19:10:16 2026

Generated:

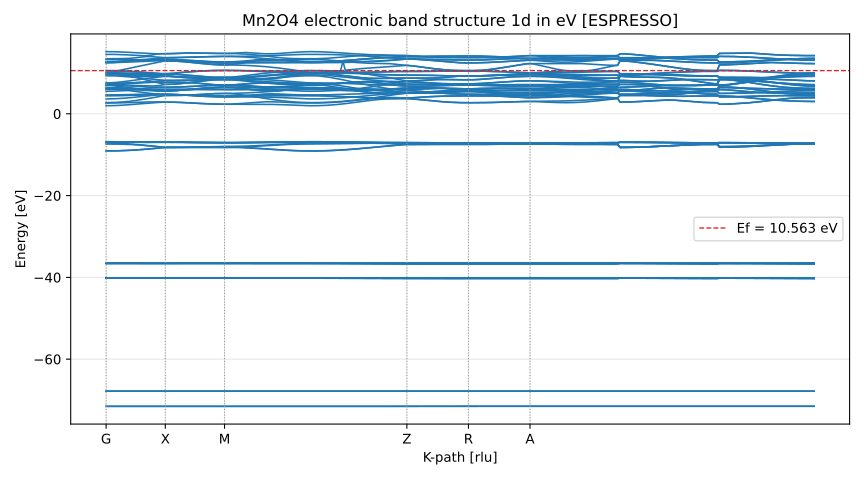
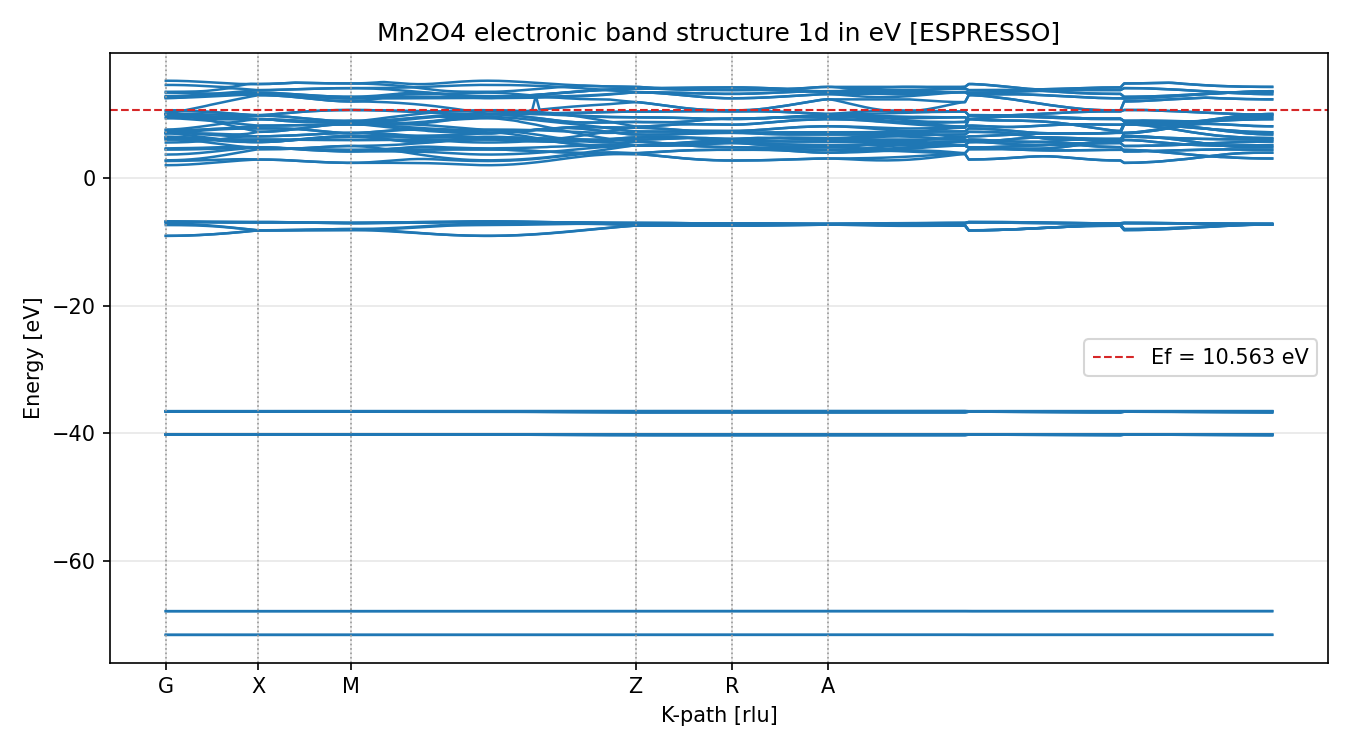
Electronic band structure along k-path on Sun Nov 1 19:14:52 2026

Generated:

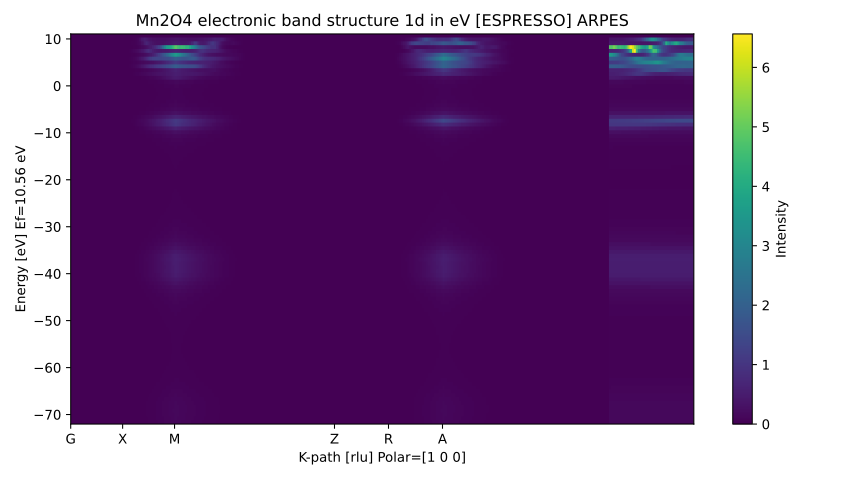
Electronic band structure in full BZ on Sun Nov 1 19:32:14 2026

Generated:

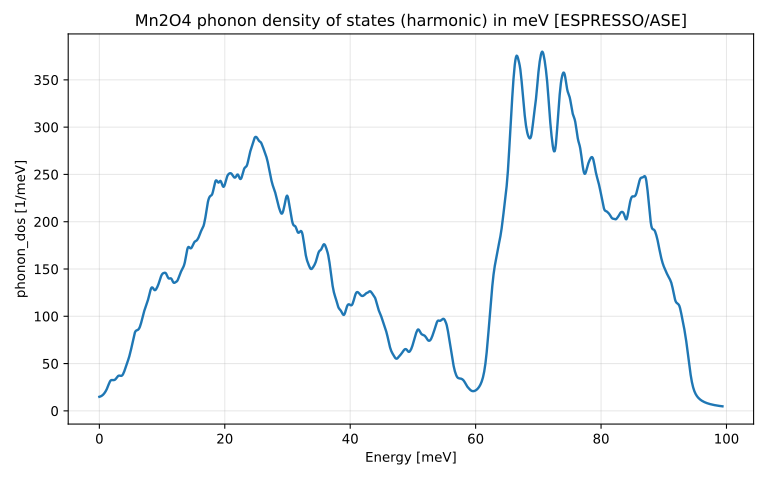
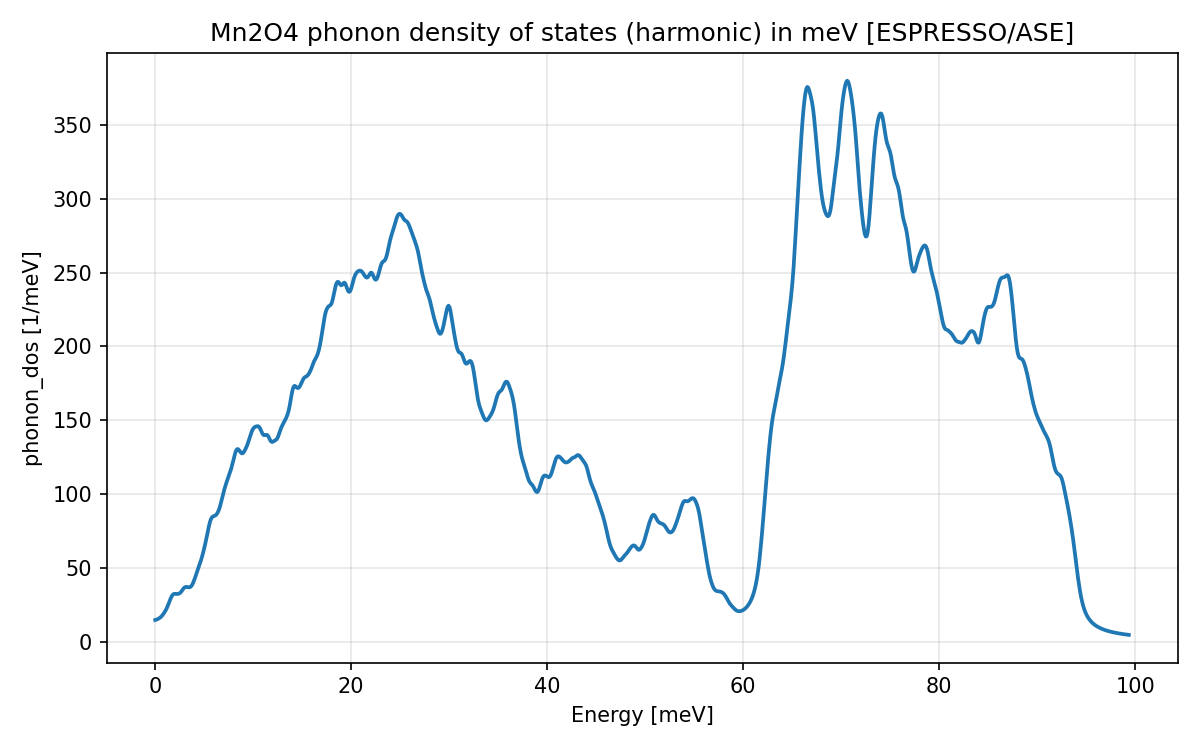
Vibrational density of states (harmonic) on Mon Nov 2 08:04:17 2026

Generated:

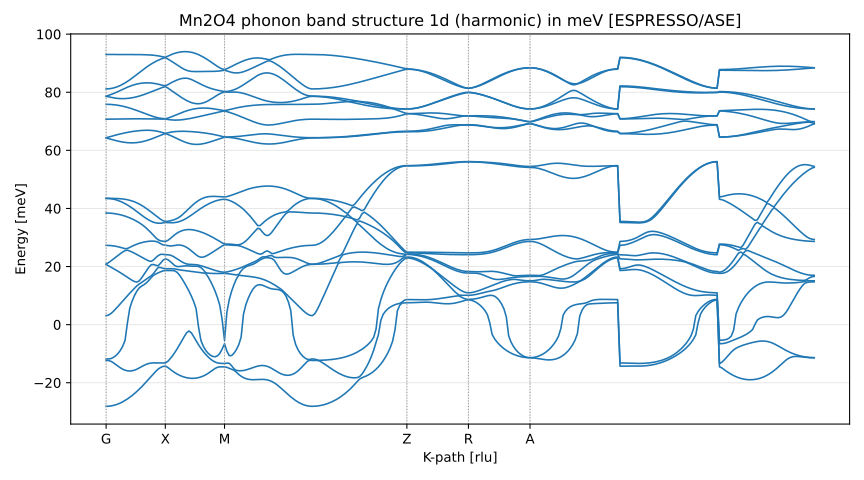
Vibrational band structure along k-path (harmonic) on Mon Nov 2 08:04:35 2026

Generated:

Vibrational band structure in full BZ (harmonic) on Mon Nov 2 08:04:52 2026

Generated:

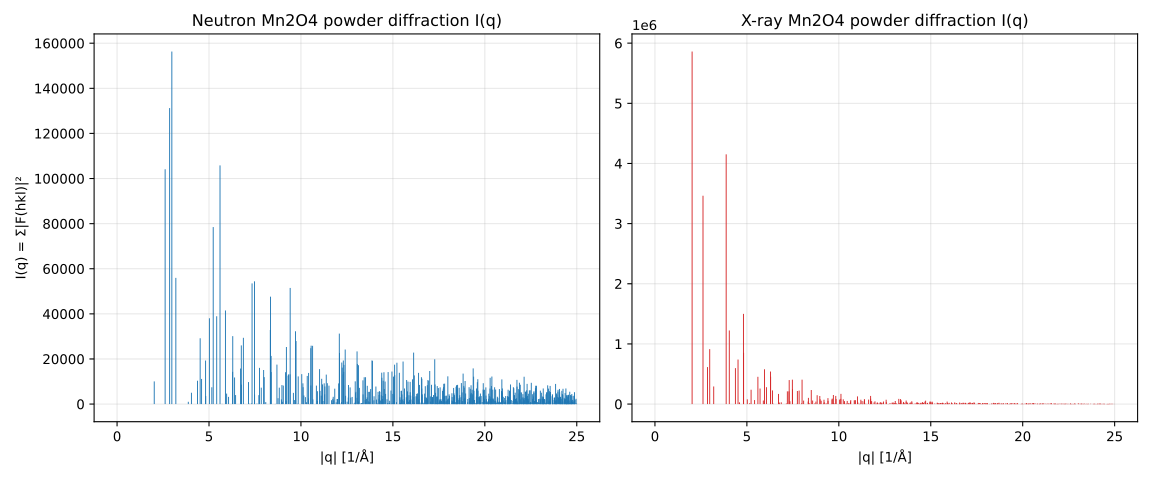
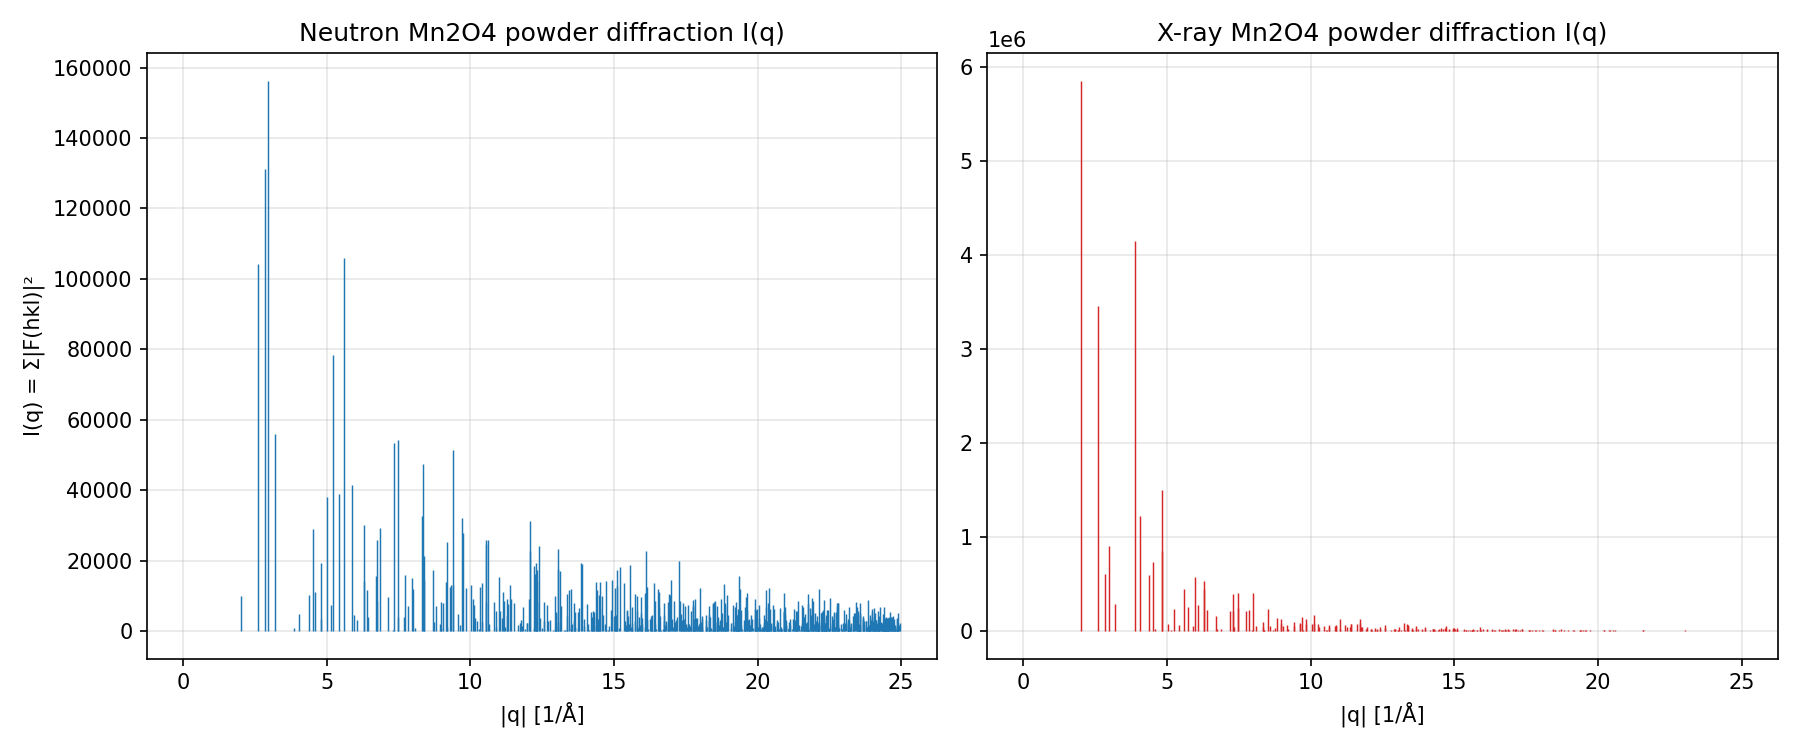
Powder diffraction I(q) (neutrons and X-rays) on Mon Nov 2 08:04:55 2026

Generated:

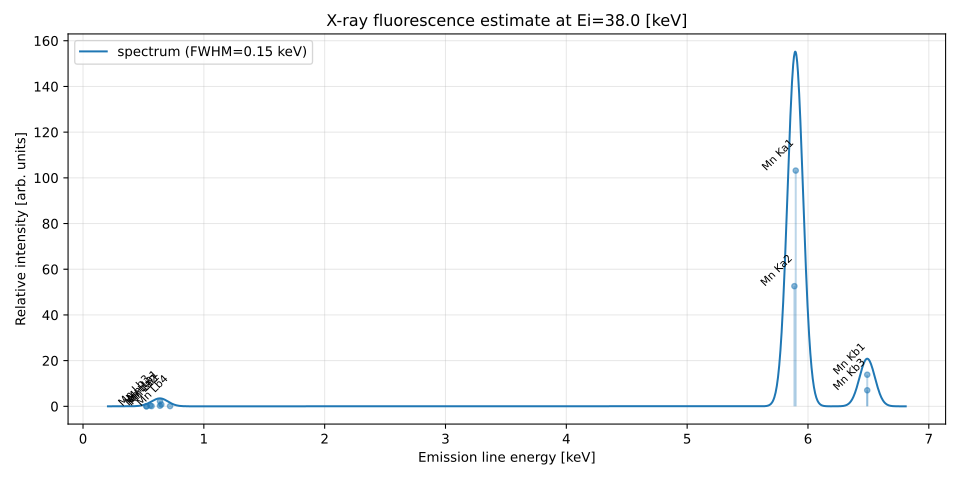
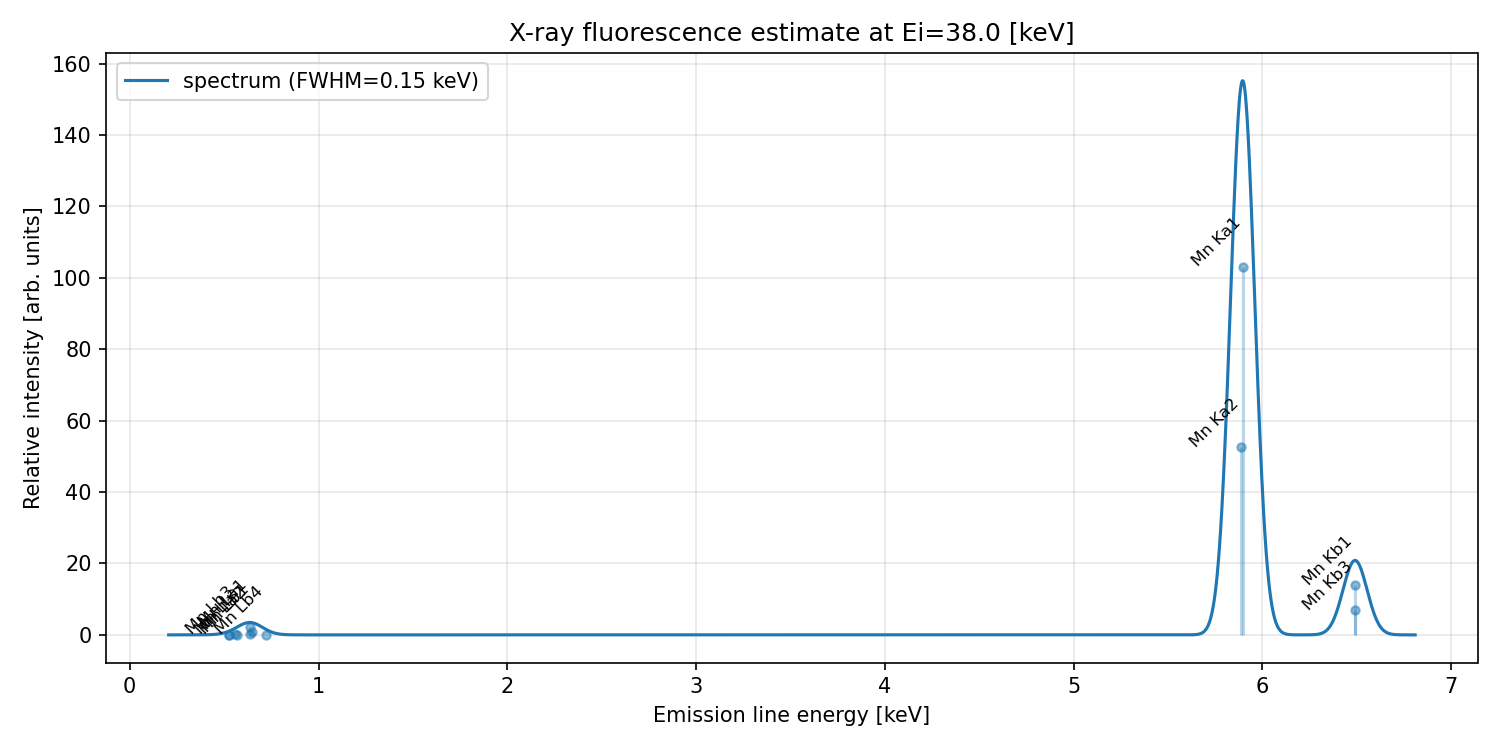
X-ray fluorescence estimate on Mon Nov 2 08:04:55 2026

Generated:

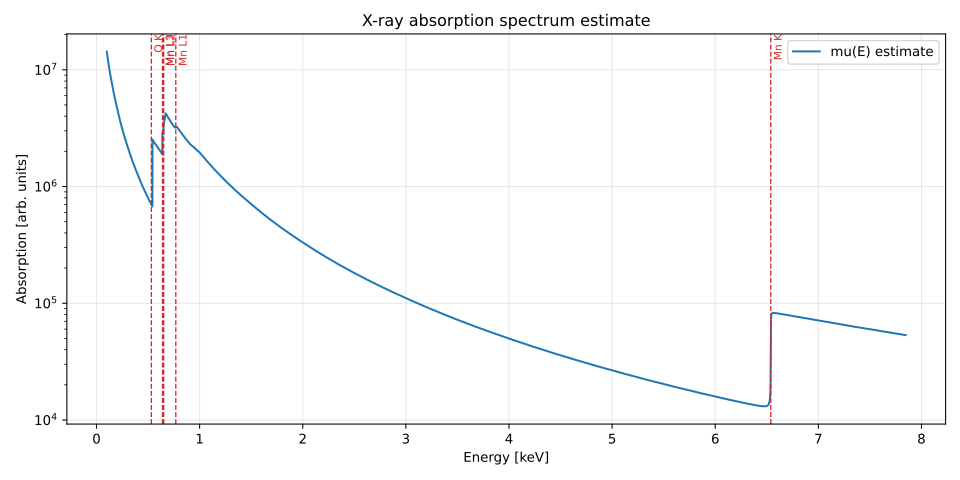
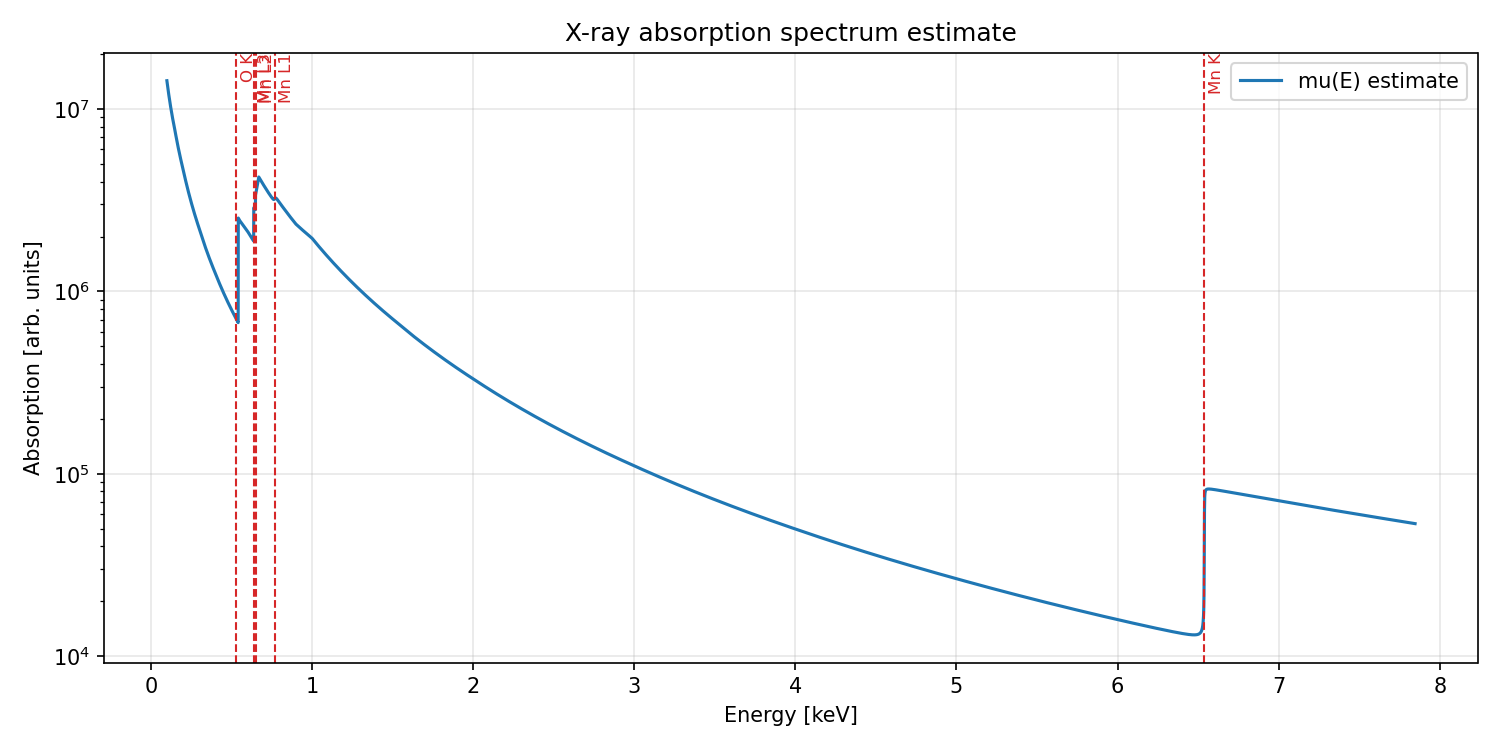
X-ray absorption spectrum estimate on Mon Nov 2 08:04:56 2026

Generated:

End of calculation on Mon Nov 2 08:05:15 2026

Generated:

The computation results are available as an HDF5 file. View it with silx view properties.h5


Duration: 46497.53339147568

End of Report

We recommend to use silx to view HDF5 and NPY files, and paraview for NRRD files.


SOLEIL(c) Synchrotron SOLEIL GRADES/EXP Project code at Gitlab